


バーチャルスクリーニング
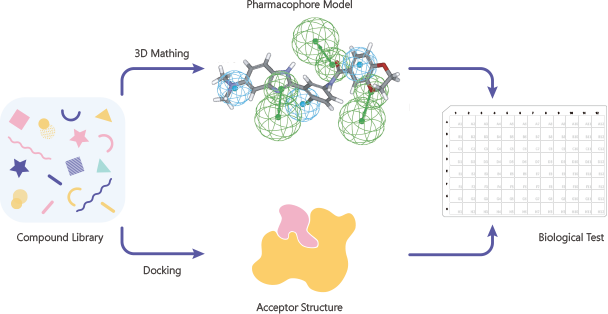
In the last decades, high-throughput screening (HTS), which refers the experimental screening of large libraries of chemicals against a biological target, plays a crucial role in the identification of new lead compounds in the early-stage drug discovery. However, HTS requires expensive equipment and facilities, and its success depends on the size of the compound library. The high cost and low hit rate associated with HTS have stimulated the development of in silico virtual screening (VS). Virtual screening is a computational technique used to search libraries of small molecules in order to identify those structures which are most likely to bind to a drug target. Nowadays, it has become a crucial step in early-stage drug discovery owing to its unique advantages over experimental HTS: drug target-relevant, competitive price and efficient.
MedChemExpress (MCE) provides high quality virtual screening service that enables researchers to identify most promising candidates. Based on the laws of quantum and molecular physics, our virtual screening services can achieve highly accurate results. Our optimized virtual screening protocol can reduce the size of chemical library to be screened experimentally, increase the likelihood to find innovative hits in a faster and less expensive manner, and mitigate the risk of failure in the lead optimization process.
Publications Citing Use of MCE Virtual Screening (Part)
- Signal Transduct Target Ther. 2025 Oct 7;10(1):333.
- Circulation. 2025 Sep 23;152(12):882-898.
- Cell Death Differ. 2025 Apr;32(4):613-631.
- Adv Sci (Weinh). 2025 Aug;12(31):e14902.
- Adv Sci (Weinh). 2025 Jul 17:e07167.
- Adv Sci (Weinh). 2025 Jul 16:e17278.
- Nat Chem Biol. 2025 Jun 2.
- Theranostics. 2025 Mar 18;15(9):4188–4211.
- Environment International 198 (2025) 109442
- Cell Death Dis. 2025 Apr 29;16(1):351.
- NPJ Biofilms Microbiomes. 2025 Mar 7;11(1):40.
- Nat Commun. 2024 Jan 2;15(1):9.
- Nat Commun. 2024 May 17;15(1):4216.
- Adv Sci (Weinh). 2024 Oct;11(39):e2402115.
- Adv Sci (Weinh). 2025 Feb;12(6):e2410525
- Theranostics. 2024 Jul 1;14(10):3984-3996.
- J Hematol Oncol. 2022 Jul 14;15(1):93.
- Theranostics. 2022 Sep 6;12(15):6509-6526.
- Cancer Lett. 2022 Oct 10:546:215851.
- J Transl Med. 2024 Feb 21;22(1):192
Types
The virtual screening methods are mainly divided into two types: structure-based virtual screening (SBVS) and ligand-based virtual screening (LBVS).
• SBVS
The general scheme of a SBVS strategy starts with processing the 3D target structural information of pharmaceutical protein interested (determined either experimentally or computationally through homology modeling) and then dock the small molecules to targeted binding sites. These docked compounds are then ranked based on their predicted binding affinity or complementarity to the binding site, as well as other criteria. Usually only a few top-ranked compounds are selected as candidates for further experimental assays. Our fast and accurate ligand docking and scoring procedures lead to efficient virtual screening.
• LBVS
In the absence of 3D structures of potential drug targets, LBVS is one of the most popular approaches for drug discovery and lead optimization. Biological data are explored in order to identify known active or inactive compounds that will be used to retrieve other potentially active molecular scaffolds for experimental evaluation. LBVS methods include approaches such as similarity and substructure searching, quantitative structure-activity relationships (QSAR), pharmacophore mapping, and machine learning.
Screening Process
• Target research
• Model building
• Preparation of small molecule compound library
• Molecular docking/pharmacophore mapping
• Scoring/ranking
• Compound selection
Advantages
• Ligand-based and structure-based virtual screening
• Super high-performance computer
• Compound database containing over 16 million purchasable compounds
• 3D pharmacophore model building
• Consideration of water and solvation effects

Compound Databases
| Cat. No. | Name | Size | Description |
|---|
MedChemExpress (MCE) virtual screening services can significantly improve the hit rates and reduce the costs of compound screening. If you have any questions, please do not hesitate to contact us via email [email protected].
バーチャルスクリーニング FAQ
01 バーチャルスクリーニング解析を依頼するために必要な情報は何ですか?
- • タンパク質名および種
- • アミノ酸配列(任意)
- • 三次元構造情報(X線結晶構造、AlphaFold 予測構造など)
- • 既知の活性化合物や結合部位情報(任意)
02 使用できるのはバーチャルライブラリー(品番末尾 V)のみですか?
03 Schrödinger、AlphaFold とは何ですか?
Schrödinger:世界的に標準的な分子シミュレーション・計算化学ソフトウェア。
AlphaFold:Google DeepMind が開発した AI タンパク質構造予測システム。アミノ酸配列から高精度で 3D 構造を予測します。MCE では AlphaFold 予測構造を用いた VS も対応しています。
04 タンパク質の立体構造情報がない場合(X線構造なし)でも VS を依頼できますか?
05 MCE はいつから VS サービスを開始したのですか?
06 MCE における VS 実績について教えてください。
07 VS の委託では HTVS / SP / XP の全モード使用が前提ですか?追加費用は発生しますか?
08 VS の結果、ヒット化合物が見つからない確率はどれくらいですか?
09 VS ライブラリーのデータセットでは、法規制化合物(毒物など)は除外されていますか?PAINS 化合物は除外されていますか?
- • 法規制化合物・既知毒性化合物:原則として除外済み
- • PAINS 化合物:ライブラリー構築時に積極的にフィルタリング さらに、ドラッグライクネス評価と組み合わせて高品質な化合物セットを提供しています。
10 VS で絞り込まれた候補群に A 化合物が含まれていた場合、A の類縁化合物も同時に候補として出てきますか?
11 データの再現性はどれくらいですか?
12 納期・価格の目安は?モデルケースはありますか?
| The Number of Compounds Screened | Lead Time | Regular VS Price (USD) | Advanced VS Price (USD) | AI Screening Price (USD) |
|---|---|---|---|---|
| 50K | 1-2 Weeks | 1,000 | 2,000 | / |
| 100K | 2-3 Weeks | 1,600 | 3,200 | 4,200 |
| 200K | 3-4 Weeks | 2,200 | 4,400 | 5,300 |
| 300K | 4 Weeks | 2,700 | 5,400 | 6,300 |
| 500K | 4-5 Weeks | 3,600 | 7,200 | 8,300 |
| 1M | 5-6 Weeks | 4,600 | 9,200 | 9,200 |
| 2M | 5-6 Weeks | 6,600 | 12,200 | 12,200 |
| 5M | 5-6 Weeks | 10,000 | 20,000 | 20,000 |
| 10M | 5-6 Weeks | 16,000 | 32,000 | 32,000 |
13 見積りから支払いまでの流れは?
- 1. 相談:ターゲット情報・要望の共有
- 2. 見積り:MCE が評価し正式な見積りを提示
- 3. 確認・契約:合意後に契約締結
- 4. プロジェクト開始:タンパク質準備、化合物準備、ドッキングなど VS 実施
- 5. 納品:化合物構造、スコア、結合様式図などを納品
14 ADME や毒性予測も可能ですか?
15 特定化合物に結合するタンパク質を探索することは可能ですか?
- • 価格:3000 USD
- • 期間:3〜4 週間 です。
16 一般的な研究室の PC(通常の CPU 搭載)で解析は可能ですか?
17 競合メーカー(Enamine)と MCE の VS の違い、立ち位置の違いは?
- Enamine:世界最大級の化合物供給企業。REAL データベースなど、数百万〜数十億規模の化合物空間を保有。強みは「化合物」。
- MCE:VS、HTS、DEL、生物学評価まで提供する一貫型の薬物探索サービス企業。200 種以上・約 2800 万化合物のライブラリーを保有し、Schrödinger ベースの専門的 VS を提供。強みは「サービスとソリューション」。また、協業により Enamine ライブラリーの利用も可能で、両者は補完関係にあります。