1545 Results for "

efficacy

" in MedChemExpress (MCE) Product Catalog:
Products (1545)

1545 Results for "efficacy" in MCE Product Catalog:

Cat. No.: HY-178032
PARP1-IN-44, an Olaparib (HY-10162) derivative, is an orally active PARP1 inhibitor (IC50 = 0.6 nM), and also inhibits PARP2 (IC50 = 1.0 nM) and PARP7 (IC50 = 7.5 nM). PARP1-IN-44 has selective antiproliferative activity against BRCA-deficient cancer cells with minimal toxicity to normal cells. PARP1-IN-44 induces G2/M phase arrest, promotes apoptosis, elevates ROS levels, disrupts mitochondrial membrane potential. PARP1-IN-44 suppresses PARylation while increasing γH2AX accumulation. PARP1-IN-44 activates the cGAS-STING pathway, upregulating IFN-β and CXCL10 expression. PARP1-IN-44 enhancing CD8+ T cell infiltration in a CT26 tumor mouse model, demonstrating robust in vivo antitumor efficacy .
loading...
    loading...
Cat. No.: HY-188120
AZD5335 is an ADC targeting FRα, with an IC50 of 10.6 nM against human FRα. AZD5335 specifically binds to FRα, mediates internalization and trafficking to lysosomal compartments, while delivering a topoisomerase I inhibitor payload that traps TOP1-DNA cleavage complexes. AZD5335 induces DNA damage response via the ATR, ATM, Chk1 and Chk2 signaling pathways, and increases the levels of phosphorylated KAP1, phosphorylated RPA, phosphorylated γH2AX, as well as PARP cleavage. AZD5335 exhibits bystander cytotoxicity against FRα-negative cells co-cultured with FRα-expressing cells. AZD5335 induces potent and durable responses and complete regression in ovarian cancer models, including models with low FRα expression and models resistant to Elahere, with enhanced efficacy when combined with Carboplatin, Bevacizumab or poly (ADP-ribose) polymerase inhibitors. AZD5335 can be used for ovarian cancer research .
loading...
    loading...
Cat. No.: HY-W720629
CAS No.: 21420-37-1
Cyanamide- 15N2 is the 15N-labeled Cyanamide (HY-Y0070). Cyanamide is a cell division and plant growth inhibitor, as well as an allelochemical derived from Vicia villosa. Cyanamide inhibits root growth and biomass accumulation in a dose-dependent manner by disrupting the formation of mitotic spindles and phragmoplast complexes, reducing the number of mitotic cells and blocking the cell cycle. The effects of Cyanamide are partially reversible after removal from low-concentration environments. Cyanamide is also a specific inhibitor of aldehyde dehydrogenase (ALDH). Although Cyanamide has no direct effect on tumor growth, it can significantly enhance the anti-tumor efficacy of Cyclophosphamide (HY-17420) at non-toxic doses by inhibiting the inactivation of Cyclophosphamide. Cyanamide enables Cyclophosphamide to exert equivalent therapeutic effects at lower doses, effectively inhibiting the growth of primary and metastatic tumors and prolonging the lifespan of tumor-bearing mice. Cyanamide is commonly used in studies related to ha-1 hepatoma and rls lymphosarcoma .
loading...
    loading...
Cat. No.: HY-164919
CAS No.: 2847102-44-5
Synonyms: XMT-2056
Calotatug ginistinag (XMT-2056) is an antibody-drug conjugate (ADC) targeting HER2, linked to a payload composed of XMT-1519 conjugate-1 (HY-148067) and STING agonist-20 (HY-148068). Calotatug ginistinag activates the STING signaling pathway in tumor cells and tumor-resident immune cells, induces the production of type I interferons and cytokines (CXCL10, IFN-β, IL-6, TNF-α, triggers innate anti-tumor immune responses, and exerts a bystander effect. Calotatug ginistinag exhibits efficacy against HER2-expressing cancer cells in co-culture with PBMCs. Calotatug ginistinag induces tumor regression and reduces systemic inflammation in various tumor models. Combination treatment with Calotatug ginistinag, Trastuzumab (HY-P9907) and T-DXd (HY-138298) yields benefits. Calotatug ginistinag can be used in studies related to HER2-expressing solid tumors such as breast cancer, gastric cancer and ovarian cancer .
loading...
    loading...
Cat. No.: HY-177578
NN3201 is a c-Kit-targeting antibody-drug conjugate (ADC) with high affinity (KD = 0.19 pM). NN3201 is composed of 4-(3-Tosyl-2-(tosylmethyl)propanoyl)benzoic acid-glu(PEG24-Me)-val-cit-NH-benzyloxyformic acid-MMAE (HY-178219) and an anti-c-Kit human monoclonal antibody NN2101 (HY-P991293). NN3201 rapidly internalizes and inhibits stem cell factor (SCF)-driven signaling, thereby delivering its payload to induce cell cycle arrest and apoptosis. NN3201 exhibits no Fc-mediated effector functions antibody-dependent cell-mediated cytotoxicity (ADCC)/complement-dependent cytotoxicity (CDC) due to reduced FcγR binding. NN3201 exhibits significant c-Kit-dependent anti-tumor efficacies in various tumor models. NN3201 can be used in small cell lung cancer (SCLC) and gastrointestinal stromal tumor (GIST) and acute myeloid leukemia (AML) research [1][2].
loading...
    loading...
Cat. No.: HY-178858
PROTAC FLT3/CHK1 Degrader-1 is a PROTAC FLT3/CHK1 degrader, with DC50 values of 5.88 nM (FLT3) and 4.17 nM (CHK1), respectively. PROTAC FLT3/CHK1 Degrader-1 can inhibit the phosphorylation of FLT3 downstream signaling effectors STAT5 (Tyr694), AKT (Ser473), and ERK (Tyr204), downregulate the protein level of c-Myc and maintain the expression of p53 protein. PROTAC FLT3/CHK1 Degrader-1 induces Apoptosis in cells. PROTAC FLT3/CHK1 Degrader-1 shows significant anti-tumor efficacy in mice bearing MV-4-11 subcutaneous xenografts. PROTAC FLT3/CHK1 Degrader-1 can be used for the study of acute myeloid leukemia (AML) .
loading...
    loading...
Cat. No.: HY-P991149
CAS No.: 2966936-23-0
Synonyms: YH32367; ABL105

Target:  

TNF Receptor

Research Areas:  

Cancer

Nesfrotamig (YH32367; ABL105) is a bispecific activator targeting HER2 and 4-1BB. The Kd values of Nesfrotamig for human HER2 and human 4-1BB are 0.48 nM and 3.36 nM, respectively. By blocking tumor cell growth signals, activating HER2-dependent local 4-1BB in tumors to maintain T cell survival, and inducing NK cell-mediated antibody-dependent cellular cytotoxicity, Nesfrotamig enhances the cytotoxicity and tumor infiltration ability of immune cells. Nesfrotamig promotes the generation of tumor-specific memory T cells, drives T cell-mediated tumor lysis, exhibits significant anti-tumor efficacy against both HER2-positive and HER2-low-expressing tumors, and shows synergistic activity when combined with anti-PD-1 antibodies. In cynomolgus monkey studies, Nesfrotamig demonstrates good safety and is suitable for research related to HER2-positive and HER2-low-expressing tumors .
loading...
    loading...
Cat. No.: HY-P992401
M802 is an anti-HER2/CD3 bispecific antibody, with a Kd of 0.578 nM for human HER2 and a Kd of 71.2 nM for human CD3. M802 inhibits the PI3K/AKT and MAPK signaling pathways, suppresses tumor cell proliferation, activates caspase-3, and promotes tumor cell apoptosis (apoptosis). M802 recruits and activates CD3-positive immune cells, mediates cytotoxicity against HER2-positive tumor cells, and induces immune cells to secrete IFN-γ, TNF-α, IL-2 and IL-6. M802 exhibits anti-tumor efficacy in mice with gastric cancer xenografts. M802 can be used in research related to HER2-positive breast cancer, HER2-positive gastric cancer and other cancers. The recommended isotype control is human IgG1 kappa (HY-P99001) .
loading...
    loading...
Cat. No.: HY-W015490R
CAS No.: 130-15-4
1,4-Naphthoquinone is an inhibitor with broad-spectrum inhibitory activity targeting DNA polymerase, NF-κB and monoamine oxidase (MAO-A/B), with antibacterial and anti-biofilm efficacy. 1,4-Naphthoquinone is a competitive inhibitor of MAO-B (Ki=1.4 μM) and a non-competitive inhibitor of MAO-A (Ki=7.7 μM). 1,4-Naphthoquinone inhibits DNA polymerase pol α, β, γ, δ, ε, λ with IC50 ranging from 5.57-128 μM. 1,4-Naphthoquinone inhibits tumor cell proliferation, induces apoptosis and necrosis, and has anti-angiogenic and anti-inflammatory activities by inducing oxidative stress, depleting glutathione (GSH), inhibiting DNA polymerase-mediated DNA synthesis and blocking NF-κB nuclear translocation. 1,4-Naphthoquinone can be used in anti-bacterial , anti-tumor and anti-inflammatory studies, including inhibition of melanoma and colon cancer cell growth and endothelial cell function, as well as LPS-induced inflammation models .
loading...
    loading...
Cat. No.: HY-L105S
867 compounds

Peptides, composed of amino acids, serve as crucial building blocks for proteins and have gained significant attention in drug development over the past decade. The advancements in production, modification, and analytical technologies have led to a surge in the potential applications of peptides in medicine. Peptides offer a number of advantages over small molecule drugs, including: greater target specificity and efficacy, more predictable metabolic profiles, easier delivery to where they are needed in the body, and fewer side effects. Peptides are increasingly appearing in all branches of medicine as components of innovative drugs, imaging agents, diagnostic agents, and other complex drugs such as peptide-drug conjugates. To date, more than 80 peptide drugs have been approved to treat a variety of diseases, including microbial infections, obesity, anti-diabetes, and cancer, as well as to develop cell targeting platforms and improve cell penetration properties.

MCE designs a unique collection of 867 peptide compounds. HY-L105S is a peptide compound library that can be provided with solution form based on HY-L105, and can be applied to peptides-based drug development.

Cat. No.: HY-L170
258 compounds

An emerging drug design method is based on the secondary binding site effect, where small molecule drugs are designed to bind to secondary binding sites on target biomolecules rather than primary orthomorphic sites. Successful potential drugs (known as allosteric modulators) will be able to bind to allosteric sites and remotely alter (or modify) the conformation of the main orthosteric binding sites of biological targets. Allosteric modulators (AMs) are ligands of proteins that act through binding sites different from natural (orthosteric) ligand sites. AMs are relatively small, more lipophilic, and more rigid compounds. The binding efficacy of AMs with their targets is often slightly lower. AMs are divided into positive AMs (PAMs) and negative AMs (NAMs). AMs are ideal drug targets because they can fine-tune receptor activity while preserving the spatial and temporal signal transduction characteristics of endogenous ligands, resulting in fewer targeted side effects, improved subtype selectivity, and better promotion of biased signal transduction than normal ligands.

MCE designs a unique collection of 258 small allosteric modulators. It is a good tool to be used for research on metabolize, cancer and other diseases.

Cat. No.: HY-L022M
3,236 compounds

New drug development is a time-consuming and high-cost process. Drug repurposing (also called drug repositioning, reprofiling or re‑tasking) offers various advantages over developing an entirely new drug for a given indication. First, the risk of failure is lower. Second, the time frame for drug development can be reduced. Third, less investment is needed. Approved drugs have identified bioactivities, good pharmacokinetic characteristics and safety which are suitable for drug repurposing.

MCE owns a unique collection of 3,236 approved compounds which have been completed extensive preclinical and clinical studies and have well-characterized bioactivities, safety and bioavailability properties. The package of this library is 96-well microplate with peelable foil seal, which makes the screening process easier and faster.

Cat. No.: HY-116115
CAS No.: 1233715-33-7
Synonyms: 17-Oxo-DPA; 17-Oxo-7(Z),10(Z),13(Z),15(E),19(Z)-DPA
Target:  

NF-κB PPAR

Research Areas:  

Inflammation/Immunology

17-Oxo-7(Z),10(Z),13(Z),15(E),19(Z)-docosapentaenoic acid (17-Oxo-DPA; 17-Oxo-7(Z),10(Z),13(Z),15(E),19(Z)-DPA) is an electrophilic oxo-derivative (EFOX) of the docosahexaenoic acid (DHA) (HY-B2167). 17-Oxo-7(Z),10(Z),13(Z),15(E),19(Z)-docosapentaenoic acid is generated during inflammation by COX-2-catalyzed mechanism in activated macrophages. 17-Oxo-7(Z),10(Z),13(Z),15(E),19(Z)-docosapentaenoic acid acts as an agonist for PPARγ and a modulator for NF-κB signaling pathway, inhibits the production of pro-inflammatory cytokines and nitric oxide, and exhibits anti-inflammatory efficacy .
loading...
    loading...
Cat. No.: HY-162275
CAS No.: 861224-48-8
Research Areas:  

Cancer

JMJD1C-IN-1 is an orally active and selective inhibitor of JMJD1C (IC50 = 0.59 μM, Kd = 1.96 μM). JMJD1C-IN-1 inhibits the binding of JMJD1C to H3K9me2 peptide substrate in the HTRF assay (IC50 = 1.47 μM). JMJD1C-IN-1 disrupts intratumoral regulatory T (Treg) cell fitness by dual mechanisms: promoting H3K9me2 accumulation to downregulate PD1 expression and reducing STAT3 demethylation to enhance STAT3 activation. JMJD1C-IN-1 demonstrates dose-dependent antitumor efficacy in multiple mouse tumor models (MCA205 fibrosarcoma, B16-F10 melanoma, LLC lung cancer, Hepa1-6 hepatocellular carcinoma, CT26 colorectal cancer). JMJD1C-IN-1 can be used for the study of tumor immunotherapy by selectively targeting intratumoral Treg cells .
loading...
    loading...
Cat. No.: HY-181906
Research Areas:  

Cancer

ZnPc-PEG2-VH032 is a VHL-pathway-dependent photodegradation targeting chimera (PDTAC) and cytotoxic agent. ZnPc-PEG2-VH032 (HY-120217) binds to the VHL ligand domain, and then specifically degrades VHL under light irradiation, a process independent of non-specific ROS-mediated protein damage. ZnPc-PEG2-VH032 uses Zinc phthalocyanine (HY-19204) as a photosensitizer, and generates ROS via type I and type II photodynamic pathways under 680 nm LED irradiation. On one hand, it targets and degrades the bound VHL protein through ROS; on the other hand, it exerts direct photodynamic cytotoxicity. Meanwhile, the degradation of VHL downregulates the phosphorylation level of CDK2/4, induces cell cycle arrest in tumor cells, further enhances the sensitivity of tumor cells to oxidative damage caused by ROS, and achieves a synergistic anti-tumor effect. ZnPc-PEG2-VH032 exerts significant in vivo efficacy in an orthotopic mouse model of non-muscle invasive bladder cancer (NMIBC) .
loading...
    loading...
Cat. No.: HY-183684
Target:  

RET VEGFR PDGFR Akt ERK

Research Areas:  

Endocrinology Cancer

RET-IN-33 is a moderately selective inhibitor of RET mutants. RET-IN-33 potently inhibits G810 mutants, with IC50 values of 4.43 nM (G810R), 3.28 nM (G810C) and 0.51 nM (G810S), respectively. RET-IN-33 also inhibits other RET mutants: V804M (IC50 0.73 nM), V804L (IC50 0.36 nM), Y806H (IC50 0.74 nM) and M918T (IC50 0.55 nM). RET-IN-33 also inhibits other kinases, with an IC50 of 1.50 nM against VEGFR2 and 1.60 nM against PDGFRα. RET-IN-33 blocks the autophosphorylation of RET mutants and the downstream SHC/AKT/ERK signaling pathway. RET-IN-33 selectively inhibits the proliferation of RET-driven cell models without affecting non-RET-dependent or normal cells. RET-IN-33 exhibits dose-dependent antitumor efficacy in RET-driven xenograft models. RET-IN-33 can be used for the research of medullary thyroid carcinoma, papillary thyroid carcinoma and non-small cell lung cancer .
loading...
    loading...
Cat. No.: HY-L238
2,383 compounds

Mongolian medicine refers to drugs used for disease prevention and treatment under the guidance of Mongolian medical theory. Originating from the vast grasslands and desert regions of northern China, Mongolian medicine developed under specific natural conditions and a nomadic lifestyle, forming a unique theoretical system and treatment methods. It emphasizes the balance between the human body and the natural environment and has demonstrated distinct efficacy in treating digestive system diseases, rheumatism, skin diseases, and various chronic conditions. For example, Mongolian medicine often uses Chinese red date for palpitations and angina pectoris, sea buckthorn to relieve coughs, reduce phlegm, promote blood circulation, and remove blood stasis, and scabious to clear lung heat and treat liver heat diseases. The application of these characteristic Mongolian medicines embodies the profound wisdom of Mongolian medical practice.

With the advancement of modern technology, research on Mongolian medicine continues to deepen. Systematic studies on the chemical composition and pharmacological effects of Mongolian medicines can provide new ideas and methods for modern drug development.

MCE has collected 2,383 characteristic natural products derived from Mongolian medicines, with 400+ plant sources of Mongolian medicine.

Cat. No.: HY-10032A
CAS No.: 1247874-19-6
Target:  

14-3-3 Protein

PF-477736 (hydrochloride) is an ATP-competitive Chk1 inhibitor (Ki 0.49 nM) that also inhibits CK2A1, PKCA, and Akt1, and is a broad-spectrum kinase inhibitor .PF-477736 induces apoptosis via PARP cleavage, caspase-3/7 activation, caspase-independent death, BAX and PUMA upregulation, H2AX phosphorylation, DNA fragmentation, and Chk1 proteasomal degradation, while inhibiting proliferation and preventing mitotic entry .PF-477736 demonstrates selective synthetic lethality in LIMD1-deficient cells, efficacy in subcutaneous xenograft models of LIMD1-deficient tumors, and anti-proliferative activity against leukemia and lymphoma cell lines .PF-477736 induces DNA double-strand breaks and activates the ATM-p53-p21 axis, and sensitizes CHK1i-insensitive neuroblastoma cells to apoptosis with ATM or DNA-PK inhibitors .PF-477736 can be used for the research of non-small cell lung cancer, leukemia, lymphoma, and neuroblastoma .
loading...
    loading...
Cat. No.: HY-W015490S
CAS No.: 26473-08-5
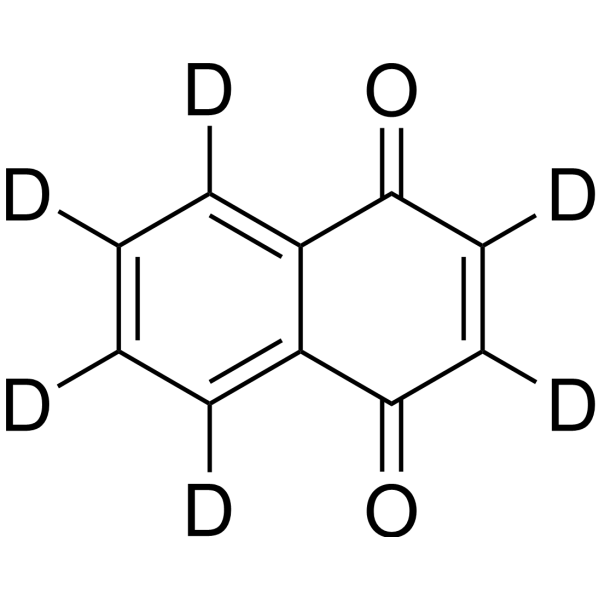
1,4-Naphthoquinone-d6 is the deuterium labeled 1,4-Naphthoquinone. 1,4-Naphthoquinone is an inhibitor with broad-spectrum inhibitory activity targeting DNA polymerase, NF-κB and monoamine oxidase (MAO-A/B), with antibacterial and anti-biofilm efficacy. 1,4-Naphthoquinone is a competitive inhibitor of MAO-B (Ki=1.4 μM) and a non-competitive inhibitor of MAO-A (Ki=7.7 μM). 1,4-Naphthoquinone inhibits DNA polymerase pol α, β, γ, δ, ε, λ with IC50 ranging from 5.57-128 μM. 1,4-Naphthoquinone inhibits tumor cell proliferation, induces apoptosis and necrosis, and has anti-angiogenic and anti-inflammatory activities by inducing oxidative stress, depleting glutathione (GSH), inhibiting DNA polymerase-mediated DNA synthesis and blocking NF-κB nuclear translocation. 1,4-Naphthoquinone can be used in anti-bacterial , anti-tumor and anti-inflammatory studies, including inhibition of melanoma and colon cancer cell growth and endothelial cell function, as well as LPS-induced inflammation models .
loading...
    loading...
Cat. No.: HY-L929
2,527 compounds

In drug discovery and development (R&D) area, target binding and druggability optimization are core processes. Among these attributes, high solubility is critical for a compound to achieve druggability, as it directly impacts the progress of drug R&D. Superior solubility ensures the rapid dissolution and uniform distribution of drug molecules in vivo, thereby enhancing bioavailability and effectively mitigating issues such as suboptimal efficacy, increased dosage requirements, or exacerbated toxic and side effects arising from insufficient solubility.

From the perspective of medicinal chemistry, high-solubility drug fragments serve as high-quality "molecular building blocks". Based on these fragments, lead compounds with potential druggability can be rapidly screened out, which significantly shortens the drug R&D cycle and reduces R&D costs. Meanwhile, the high-solubility drug fragment library can provide diverse options for drug development in different therapeutic areas, offer solutions for the solubility defects of existing clinical drugs, and facilitate the development of novel, highly effective targeted drugs with higher bioavailability and better safety profiles.

MCE has collected and compiled 2,527 experimentally validated small-molecule fragments with high solubility. These fragments can be directly used for drug molecular design, providing high-quality pre-validated solubility fragments that significantly improve the efficiency of lead compound screening and accelerate the progress of drug R&D.