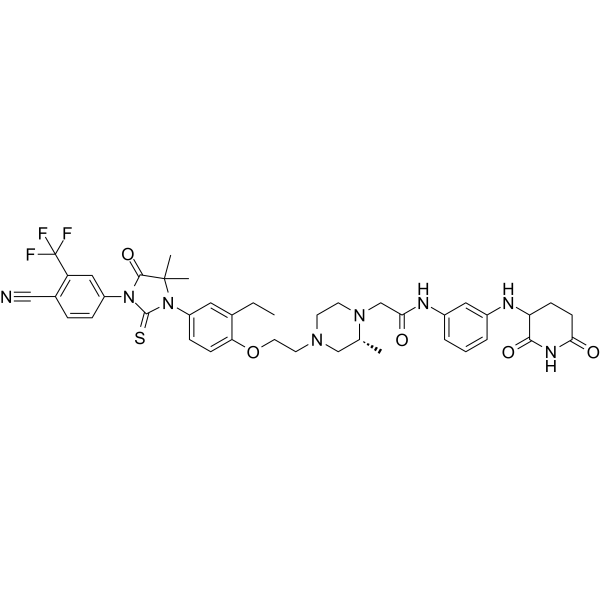
380 Results for "

mutant selective

" in MedChemExpress (MCE) Product Catalog:
Products (380)

380 Results for "mutant selective" in MCE Product Catalog:

4
4 Cited Publications
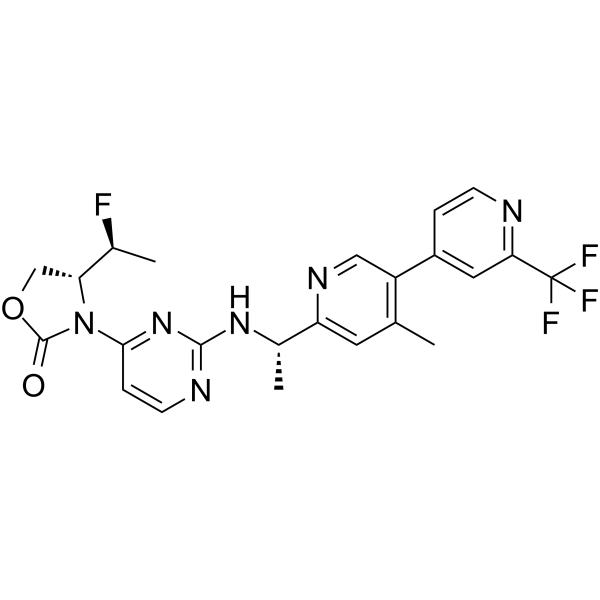
Cat. No.: HY-104036
CAS No.: 1628805-46-8
Purity:  98.75%
Research Areas:  

Cancer

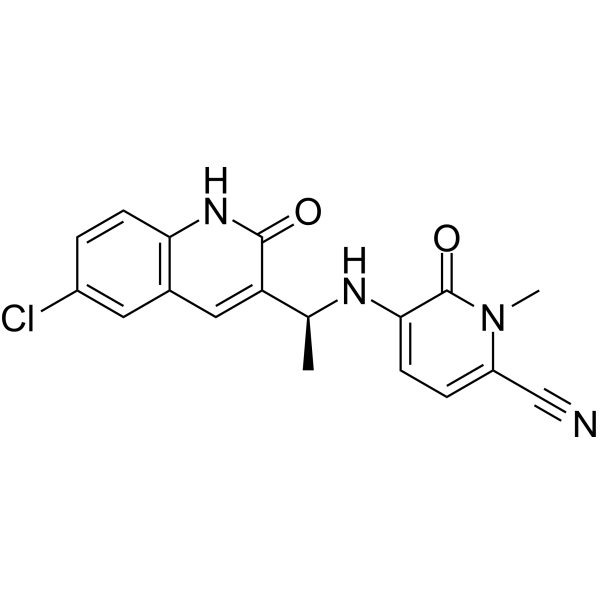
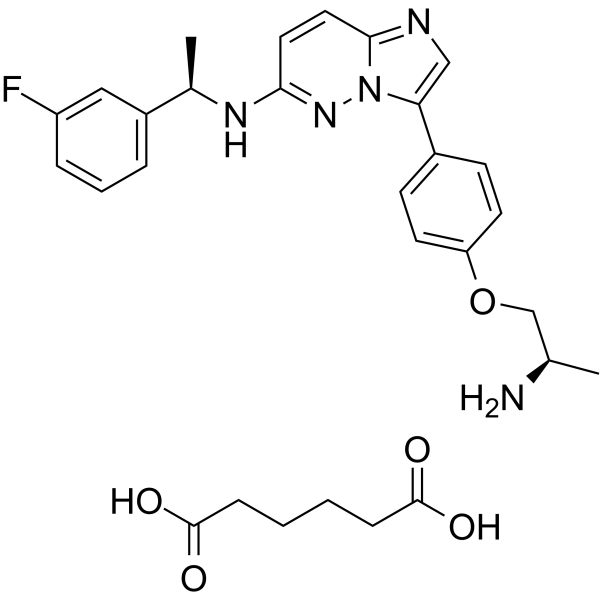
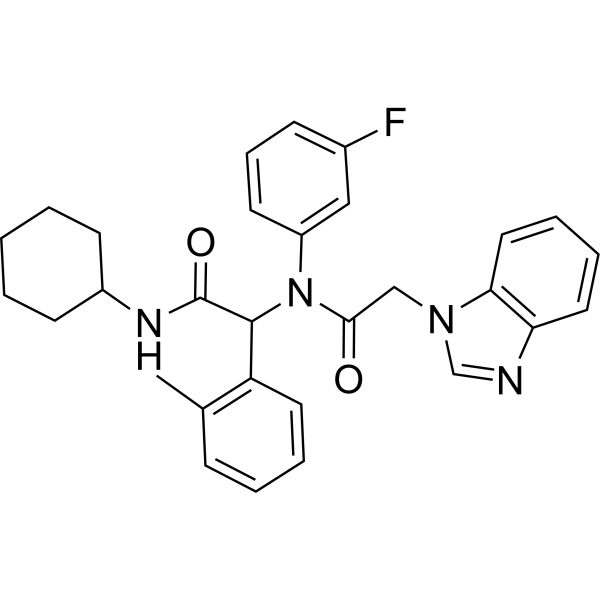
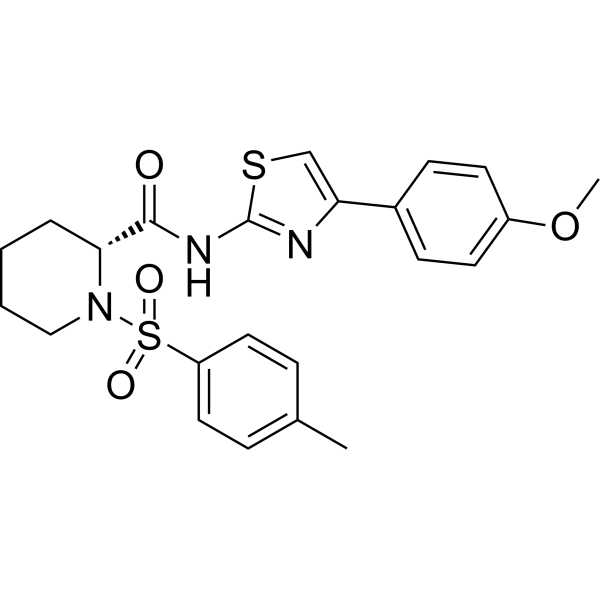
IDH-305 is an orally available, mutant-selective and brain-penetrant IDH1 inhibitor that targets IDH1 (R132) mutation. IDH-305 exhibits greater than 200 fold selectivity for mutant IDH1 isoforms vs. WT (IC50= 27 nM (IDH1 R132H), 28 nM (IDH1 R132C), 6.14 µM (IDH1 WT)) .
loading...
    loading...
4
4 Cited Publications
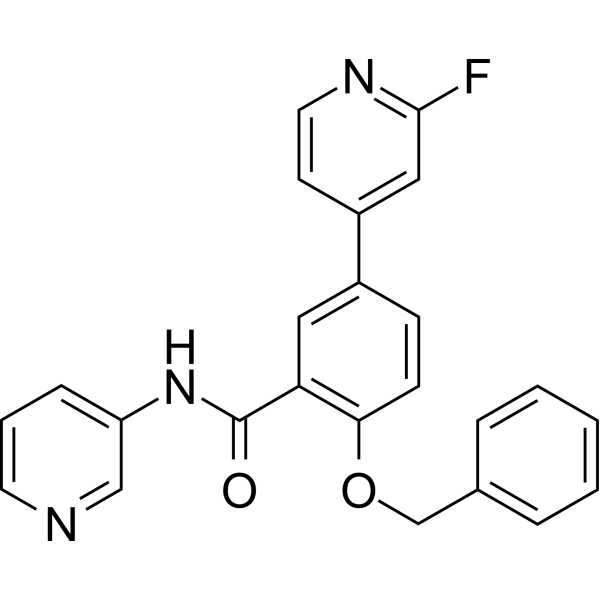
Cat. No.: HY-13237
CAS No.: 1285515-21-0
Purity:  99.82%
Target:  

LRRK2 Autophagy Mitophagy

Research Areas:  

Neurological Disease Cancer

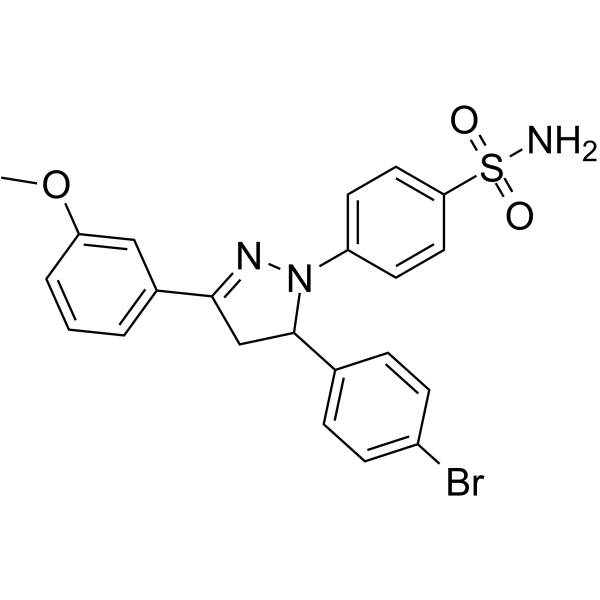
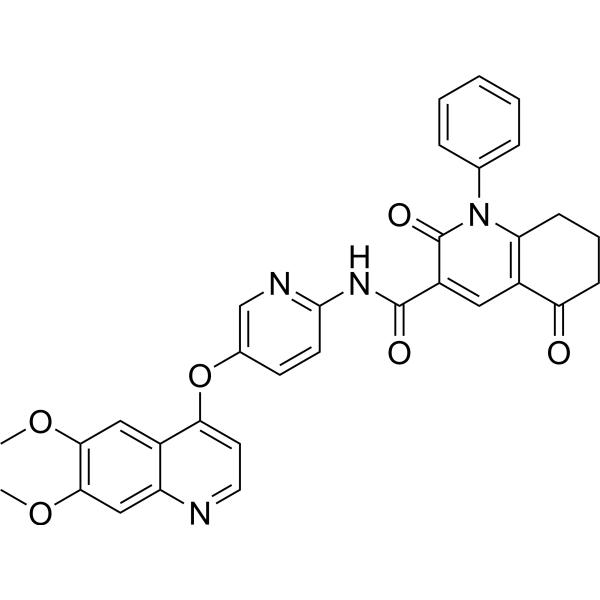
GSK2578215A is a potent and highly selective LRRK2 inhibitor, which exhibits IC50s of around 10 nM against both wild-type LRRK2 and the G2019S mutant.
loading...
    loading...
4
4 Cited Publications
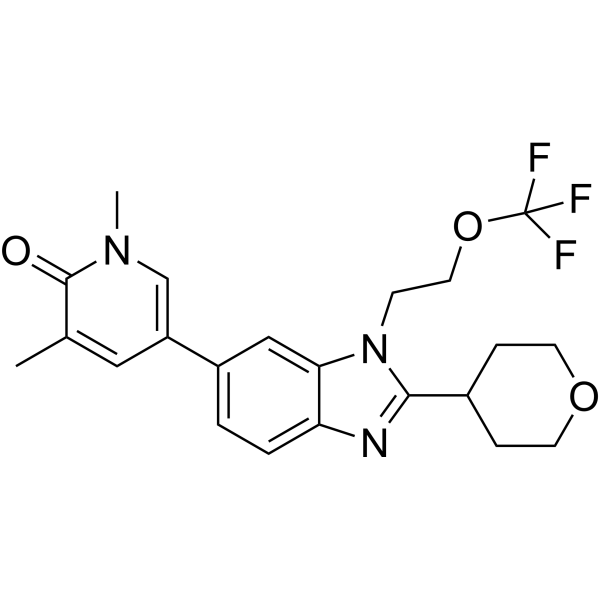
Cat. No.: HY-136938
CAS No.: 2081072-29-7
Purity:  99.63%
Synonyms: EP31670
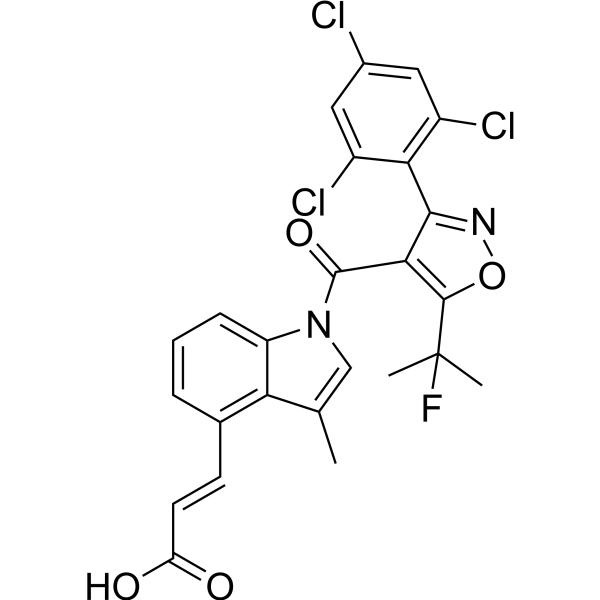
NEO2734 (EP31670) is an orally active dual p300/CBP and BET bromodomain selective inhibitor, with IC50 values of <30 nM for both p300/CBP and BET bromodomains . NEO2734 is active in SPOP mutant and wild-type prostate cancer .
loading...
    loading...
4
4 Cited Publications
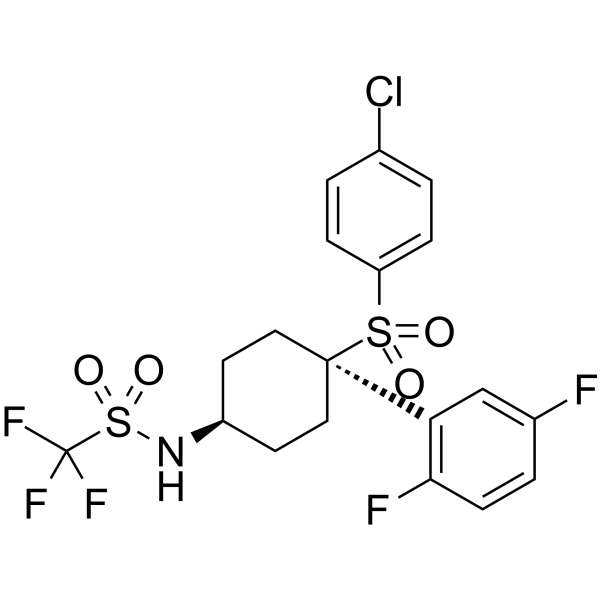
Cat. No.: HY-14174
CAS No.: 677772-84-8
Purity:  99.51%
Target:  

γ-secretase

Research Areas:  

Neurological Disease Cancer

MRK-560, a chemical probe, is an orally active, brain barrier-penetrated γ-Secretase inhibitor, reducing Aβ peptide in rat brain and cerebrospinal fluid. MRK-560 decreases mutant NOTCH1 processing by selectively inhibiting PSEN1. MRK-560 can be used in studies of Alzheimer's disease and T-cell acute lymphoblastic leukaemia (T-ALL) .
loading...
    loading...
4
4 Cited Publications
Cat. No.: HY-113137
CAS No.: 2140-67-2
Purity:  99.40%
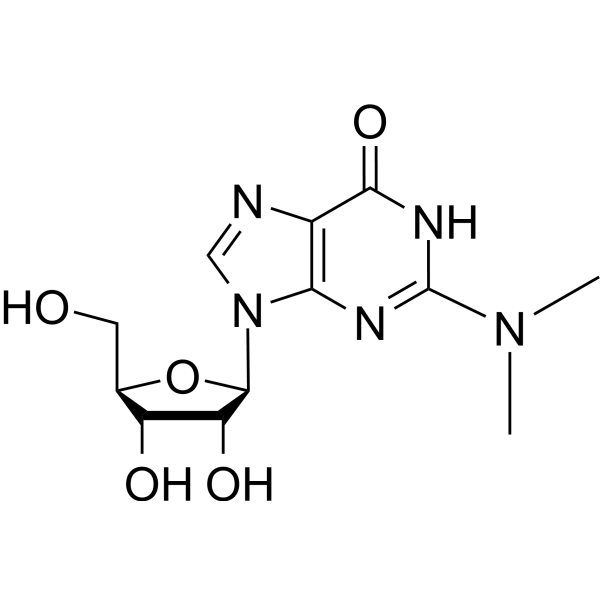
N2,N2-Dimethylguanosine is a methylated modified nucleoside present in RNA and serves as a structural modification component of tRNA. N2,N2-Dimethylguanosine inhibits reverse transcriptase-mediated cDNA synthesis and is one of the key modifications affecting sequencing efficiency in high-throughput RNA sequencing. N2,N2-Dimethylguanosine can be selectively demethylated at one methyl group by AlkB mutant enzymes (such as D135S/L118V) and converted to N2-methylguanosine, thereby reducing the inhibition of reverse transcription .
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-18997
CAS No.: 1393465-84-3
Synonyms: PB04
Target:  

Raf

Research Areas:  

Cancer

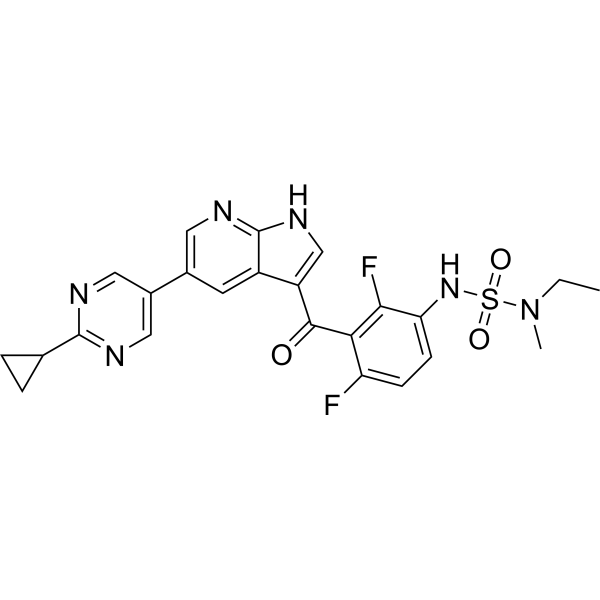
PLX7904 is a potent and selective BRAF inhibitor, with IC50 of appr 5 nM against BRAF V600E in mutant RAS expressing cells.
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-142161
CAS No.: 3007772-60-0
Purity:  99.46%
Target:  

MAGL

Research Areas:  

Cancer

ABD957 is a potent and selective covalent inhibitor of the ABHD17 family of depalmitoylases, with an IC50 of 0.21 μM for ABHD17B. ABD957 can block N-Ras signaling and the growth of NRAS-mutant AML cells .
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-114226
CAS No.: 1887014-12-1
Purity:  99.16%
Synonyms: FT-2102
Olutasidenib (FT-2102) is a highly potent, orally active, brain penetrant and selective inhibitor of mutant Isocitrate dehydrogenase 1 (IDH1), with IC50 values of 21.2 nM and 114 nM for IDH1- R132H and IDH1- R132C, respectively . Olutasidenib (FT-2102) is under the study in the treatment of acute myeloid leukemia (AML) or myelodysplastic syndrome (MDS) .
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-131003
CAS No.: 1505515-69-4
Purity:  99.78%
Synonyms: DS-6051b; AB-106; IBI-344
Target:  

ROS Kinase

Research Areas:  

Cancer

Taletrectinib (DS-6051b) is a potent, orally active, and next-generation selective ROS1/NTRK inhibitor. Taletrectinib potently inhibits recombinant ROS1, NTRK1, NTRK2, and NTRK3 with IC50s of 0.207, 0.622, 2.28, and 0.98 nM, respectively. Taletrectinib also inhibits ROS1 G2032R and other Crizotinib-resistant ROS1 mutants .
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-B0984
CAS No.: 13636-18-5
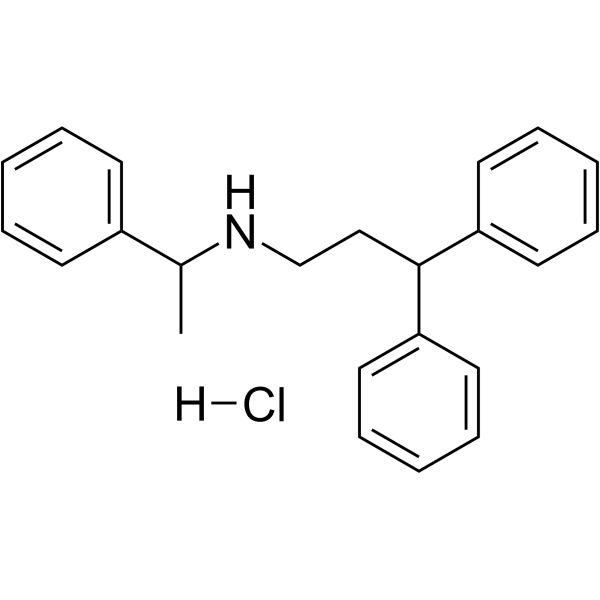
Fendiline hydrochloride, a diphenylalkylamine type of antianginal agent, is an L-type calcium channel blocker (IC50 of 17 µM). Fendiline hydrochloride is also a selective K-Ras inhibitor, and has no effect on H-Ras and N-Ras. Fendiline hydrochloride inhibits K-Ras plasma membrane localization (IC50 of 9.64 μM), inhibits K-Ras signal output and blocks the proliferation of pancreatic, colon, lung, and endometrial cancer cell lines expressing oncogenic mutant K-Ras. Fendiline hydrochloride is a STING agonist and is able to inhibit the growth of multiple refractory cold tumors (MC38, CT26 and B16F10) .
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-B0984A
CAS No.: 13042-18-7
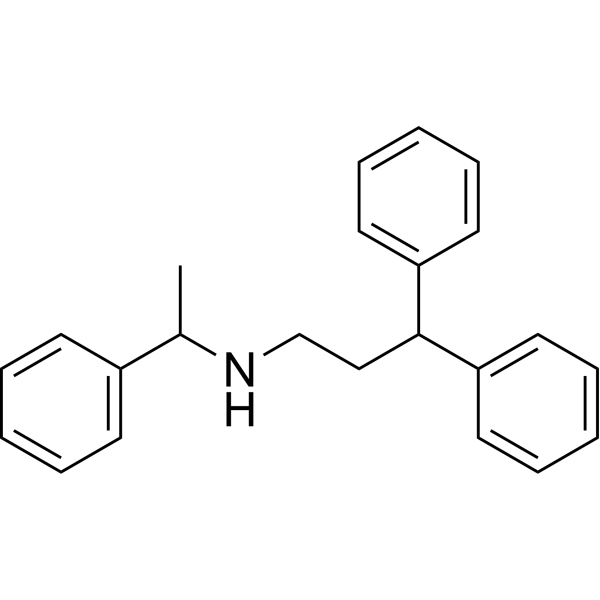
Fendiline, a diphenylalkylamine type of antianginal agent, is an L-type calcium channel blocker (IC50 of 17 µM). Fendiline is also a selective K-Ras inhibitor, and has no effect on H-Ras and N-Ras. Fendiline inhibits K-Ras plasma membrane localization (IC50 of 9.64 μM), inhibits K-Ras signal output and blocks the proliferation of pancreatic, colon, lung, and endometrial cancer cell lines expressing oncogenic mutant K-Ras. Fendiline is a STING agonist and is able to inhibit the growth of multiple refractory cold tumors (MC38, CT26 and B16F10) .
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-15466
CAS No.: 1007207-67-1
Purity:  99.01%
Synonyms: CH5132799
Target:  

PI3K Apoptosis mTOR Akt

Research Areas:  

Cancer

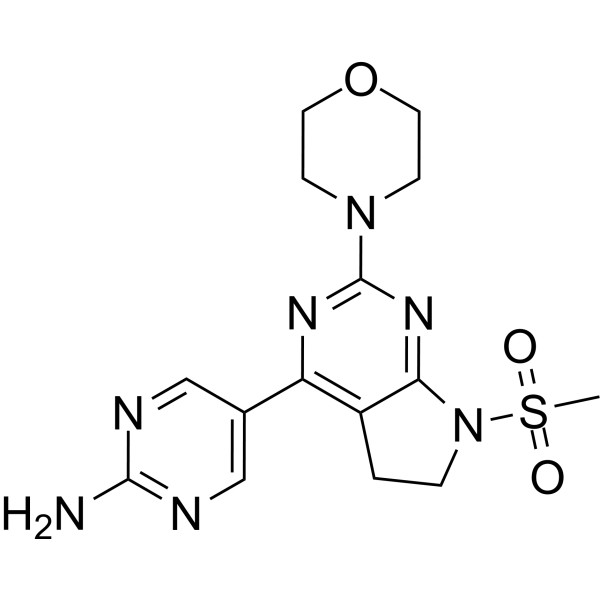
Izorlisib (CH5132799) is an orally active, selective Class I PI3K inhibitor with an IC50 value of 14 nM against PI3Kα. Izorlisib specifically inhibits Class I PI3K (particularly PI3Kα and its mutants) and blocks the PI3K/Akt/mTOR pathway; this leads to cell cycle arrest at the G1 phase and apoptosis without triggering feedback activation of Akt by competitively binding to the ATP-binding site of PI3K. Izorlisib can be used in research on cancers harboring PIK3CA mutations or PTEN loss (such as breast, ovarian, prostate, and endometrial cancers) .
loading...
    loading...
3
3 Cited Publications
Cat. No.: HY-158101
CAS No.: 2446928-30-7
Purity:  99.92%
Synonyms: CC-94676
Research Areas:  

Cancer

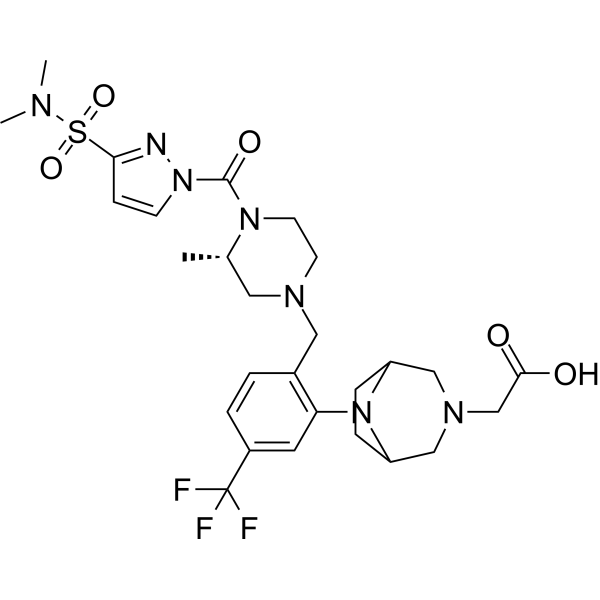
BMS-986365 (CC-94676) is an orally active and selective targeted androgen receptor (AR) PROTAC degrader (DC50 of 10-40 nM). BMS-986365 is capable of inducing cereblon (CRBN) E3 ligase-dependent ubiquitination and degradation of the androgen receptor (AR), as well as various AR mutants. BMS-986365 shows no degradation of the close AR family members estrogen receptor (ER), progesterone receptor (PR), and glucocorticoid receptor (GR). BMS-986365 shows significant in vivo potency, degrading AR, inhibiting AR signaling, and restricting tumor growth in animal models of advanced prostate cancer. BMS-986365 can be used for the study of metastatic castration-resistant prostate cancer (mCRPC) .
loading...
    loading...
2
2 Cited Publications
Cat. No.: HY-12475
CAS No.: 1355326-21-4
Purity:  99.43%
Synonyms: Agios 135
Research Areas:  

Cancer

Mutant IDH1-IN-1 (Agios 135) is a selective mutant IDH1 inhibitor with activity against both IDH1 R132H and IDH1 R132C. Mutant IDH1-IN-1 reduces the level of 2-hydroxyglutarate (2-HG), stabilizes mutant IDH1 protein, and restores blocked monocyte differentiation. Mutant IDH1-IN-1 can be used in tumor-related research .
loading...
    loading...
2
2 Cited Publications
Cat. No.: HY-136379
CAS No.: 1222513-26-9
Purity:  99.84%
Synonyms: Cdc42-IN-1
Target:  

Ras

Research Areas:  

Cancer

CID44216842 (Cdc42-IN-1) is a potent Cdc42-selective guanine nucleotide binding lead inhibitor. The EC50s for Cdc42 WT and Cdc42Q61L mutant are 1.0 and 1.2 μM in GTP binding assay, respectively. The EC50s for Cdc42 WT and Cdc42Q61L mutant are 0.3 and 0.5 μM in GDP binding assay, respectively. Use as a molecular probe .
loading...
    loading...
2
2 Cited Publications
Cat. No.: HY-145594
CAS No.: 1898206-17-1
Synonyms: AB-291; DS-1001
Research Areas:  

Cancer

Safusidenib (AB-291; DS-1001) is an orally bioavailable, selective mutant IDH1 inhibitor. Safusidenib strongly inhibits mutant IDH1 but not wild-type IDH1. Safusidenib impairs tumor activity in chondrosarcoma . Safusidenib exhibits activity against IDH1R132H, and IDH1R132C with IC50s of 15, and 130 nM in assays without any preincubation, respectively .
loading...
    loading...
2
2 Cited Publications
Cat. No.: HY-12343
CAS No.: 1401242-74-7
Purity:  98.46%
Synonyms: CID-53347902
Target:  

Potassium Channel

Research Areas:  

Cardiovascular Disease

ML277 (CID-53347902) is a potent and selective activator of K(v)7.1 (KCNQ1) potassium channel activator (EC50=270 nM), rescues function of pathophysiologically important mutant channel complexes in human induced pluripotent stem cell-derived cardiomyocytes .
loading...
    loading...
2
2 Cited Publications
Cat. No.: HY-114358
CAS No.: 1646839-59-9
Purity:  98.59%
Synonyms: ONO-7475
Research Areas:  

Cancer

Tamnorzatinib (ONO-7475) is a potent, selective, and orally active Axl/Mer inhibitor with IC50 values of 0.7 nM and 1.0 nM, respectively. Tamnorzatinib sensitizes AXL-overexpressing EGFR-mutant NSCLC cells to the EGFR-TKIs, suppresses the emergence and maintenance of tolerant cells. Tamnorzatinib combines with Osimertinib (HY-15772) provides a bright promise for the study of EGFR-mutated non-small cell lung cancer (NSCLC).
loading...
    loading...
2
2 Cited Publications
Cat. No.: HY-137458A
CAS No.: 1416775-08-0
Purity:  99.13%
Synonyms: ARQ 751 trihydrochloride
Target:  

Akt

Research Areas:  

Cancer

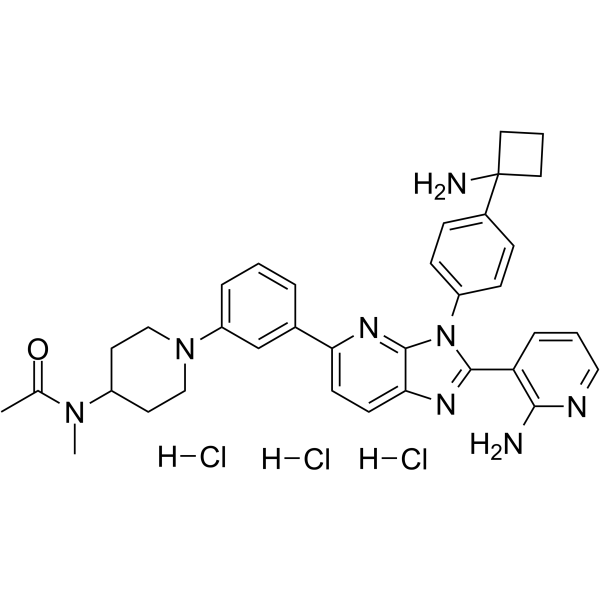
Vevorisertib (ARQ 751) trihydrochloride is a selective, allosteric, pan-AKT and AKT1-E17K mutant inhibitors. Vevorisertib trihydrochloride potently inhibit phosphorylation of AKT. Vevorisertib trihydrochloride has Kd values of 1.2 nM and 8.6 nM for AKT1 and AKT1-E17K, respectively. Vevorisertib trihydrochloride has IC50 values of 0.55, 0.81, and 1.3 nM for AKT1, AKT2, and AKT3, respectively. Vevorisertib trihydrochloride can be used for the research of cancer .
loading...
    loading...
2
2 Cited Publications
Cat. No.: HY-137516
CAS No.: 2502156-03-6
Purity:  98.16%
Research Areas:  

Cancer

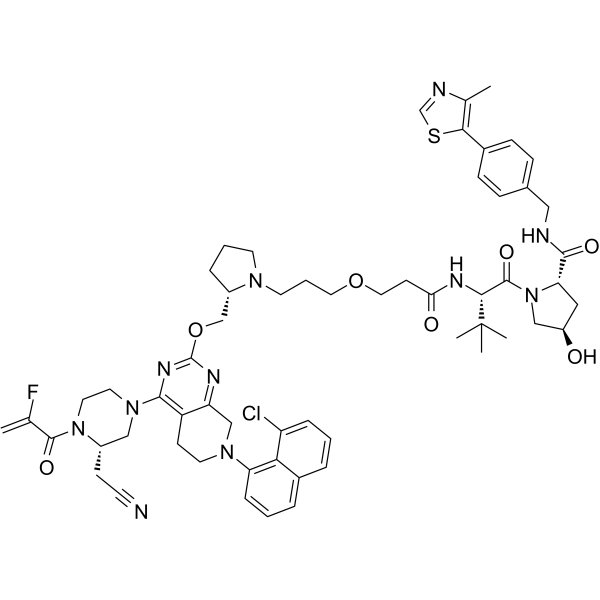
LC-2 is an orally active PROTAC-class degrader targeting KRAS G12C and a MAPK inhibitor. LC-2 shows higher selectivity for KRAS G12C than wild-type KRAS and other KRAS mutants. LC-2 covalently binds to KRAS G12C via the MRTX849 (HY-130149) warhead, recruits the VHL E3 ligase to form a ternary complex, and induces ubiquitination and degradation of KRAS G12C. Meanwhile, LC-2 inhibits the MAPK signaling pathway, reduces the phosphorylation levels of CRAF and AKT, thereby inducing apoptosis and suppressing cancer cell and tumor growth. LC-2 can be used for the research of KRAS G12C-positive cancers, including non-small cell lung cancer and colorectal cancer .
loading...
    loading...

