



PT3

Based on 1 Customer Validation
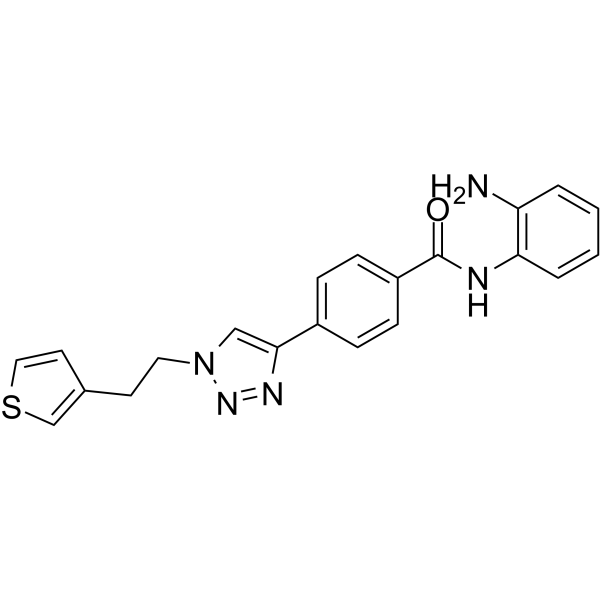
PT3 is a selective inhibitor of HDAC3 with an IC50 value of 0.25 μM. PT3 exhibits good brain penetration ability and bioavailability upon oral administration. PT3 can be used in the research of Alzheimer’s disease.
For research use only. We do not sell to patients.
- Purity : 99.94%
- CAS No.: 2710273-81-5
- Formula: C18H17N5O
- Molecular Weight:319.36
-
Storage:Powder -20°C, 3 years , 4°C, 2 years ; In solvent -80°C, 6 months , -20°C, 1 month


To place orders, for customer services and technical support, please contact: MedChemExpress USA
Tel: 609-228-6898 E-mail: [email protected] [email protected]


-
 Biological Activity
-
Chemical Information
-
Solvent & Solubility
- Protocols
- Purity & Documentation
- References
-
 Help & FAQs



Biological Activity
Description
IC50 & Target
|
HDAC2 2.08 μM (IC50) |
HDAC1 1.05 μM (IC50) |
HDAC8 15 μM (IC50) |


Chemical Information
-
CAS No. 2710273-81-5
-
Appearance Solid
-
Molecular Weight 319.36
-
Formula C18H17N5O
-
Color White to off-white
-
SMILES
O=C(C1=CN=C(NCC2=CC=CC=C2)C=N1)NC3=C(N)C=CC=C3
-
Shipping
Room temperature in continental US; may vary elsewhere.
-
Storage
Powder -20°C 3 years 4°C 2 years In solvent -80°C 6 months -20°C 1 month


Solvent & Solubility
In Vitro:
DMSO : 100 mg/mL (313.13 mM; Need ultrasonic; Hygroscopic DMSO has a significant impact on the solubility of product, please use newly opened DMSO)
Please refer to the solubility information to select the appropriate solvent. Once prepared, please aliquot and store the solution to prevent product inactivation from repeated freeze-thaw cycles.
Storage method and period of stock solution: -80°C, 6 months; -20°C, 1 month. When stored at -80°C, please use it within 6 months. When stored at -20°C, please use it within 1 month.
Concentration (start) × Volume (start) = Concentration (final) × Volume (final)

Protocols
-
How to Select the Route of Administration for Mammals
Route-of-administration selection in mammals is a pharmacokinetic, pharmacodynamic, formulation, animal-welfare, and translational decision, not a default technical choice. The selected route should match the study goal: intravenous dosing is most useful when complete systemic exposure and rapid onset are required, oral dosing is most translational for orally intended medicines but is affected by absorption and first-pass metabolism, subcutaneous or intramuscular dosing can provide slower systemic exposure, and intraperitoneal dosing can be useful in rodent proof-of-concept studies but may have limited clinical translation. Published route-comparison studies show that the same compound can produce different exposure, onset, bioavailability, tissue distribution, and tolerability depending on route; therefore, route choice should be supported by pilot pharmacokinetic or pharmacodynamic evidence when the literature is insufficient. Unresolved questions include how to standardize route sel
-
Cell Cytotoxicity Assay
Cytotoxicity assays are usually based on the assessment of cell membrane damage, which can also be indirectly detected by measuring cell viability. Detection methods include MTT assay, CKK-8 assay, LDH assay and ATP assay, etc.
-
Protocol for Pharmacokinetic Study
Pharmacokinetic studies quantify how an organism handles a drug over time through absorption, distribution, metabolism, and excretion, and the core experimental readout is the concentration-time profile of parent drug and, when relevant, metabolites in biological matrices such as plasma, whole blood, urine, bile, or tissue. Pharmacokinetic analysis links dose, route, exposure, clearance, half-life, distribution, bioavailability, and systemic exposure to drug efficacy and toxicity hypotheses rather than measuring a signaling pathway directly. The literature links pharmacokinetics to drug-development phenotypes by showing that drug metabolism and pharmacokinetics influence compound progression, exposure-response interpretation, safety margins, dosing strategy, and failure risk during discovery and development. DMPK science contributes to compound optimization by integrating physicochemical properties, in vitro metabolism, transporter behavior, in vivo exposure, and pharmacodynamic contex


Purity & Documentation
-
Data Sheet (270 KB)
-
SDS (251 KB)
- English - EN (251 KB)
- Français - FR (251 KB)
- Deutsch - DE (251 KB)
- Norwegian - NO (251 KB)
- Español - ES (251 KB)
- Swedish - SV (251 KB)
- Italian - IT (251 KB)
- Korean - KR (251 KB)
- Portuguese - PT (251 KB)
-
Handling Instructions (2659 KB)
References


Complete Stock Solution Preparation Table
Please refer to the solubility information to select the appropriate solvent. Once prepared, please aliquot and store the solution to prevent product inactivation from repeated freeze-thaw cycles.
Storage method and period of stock solution: -80°C, 6 months; -20°C, 1 month. When stored at -80°C, please use it within 6 months. When stored at -20°C, please use it within 1 month.
| Optional Solvent | Concentration Solvent Mass | 1 mg | 5 mg | 10 mg | 25 mg |
|---|---|---|---|---|---|
| DMSO | 1 mM | 3.1313 mL | 15.6563 mL | 31.3126 mL | 78.2816 mL |
| 5 mM | 0.6263 mL | 3.1313 mL | 6.2625 mL | 15.6563 mL | |
| 10 mM | 0.3131 mL | 1.5656 mL | 3.1313 mL | 7.8282 mL | |
| 15 mM | 0.2088 mL | 1.0438 mL | 2.0875 mL | 5.2188 mL | |
| 20 mM | 0.1566 mL | 0.7828 mL | 1.5656 mL | 3.9141 mL | |
| 25 mM | 0.1253 mL | 0.6263 mL | 1.2525 mL | 3.1313 mL | |
| 30 mM | 0.1044 mL | 0.5219 mL | 1.0438 mL | 2.6094 mL | |
| 40 mM | 0.0783 mL | 0.3914 mL | 0.7828 mL | 1.9570 mL | |
| 50 mM | 0.0626 mL | 0.3131 mL | 0.6263 mL | 1.5656 mL | |
| 60 mM | 0.0522 mL | 0.2609 mL | 0.5219 mL | 1.3047 mL | |
| 80 mM | 0.0391 mL | 0.1957 mL | 0.3914 mL | 0.9785 mL | |
| 100 mM | 0.0313 mL | 0.1566 mL | 0.3131 mL | 0.7828 mL |


PT3 Related Classifications
HY-164782 Related Classifications
 Powered by Bioz